Skip to content
Specialist component specification

Composite profile components for medical support systems

Define the structural blank, finished surfaces and interfaces required by the device manufacturer. Medical suitability must be established for the intended component configuration and supply scope before production release.

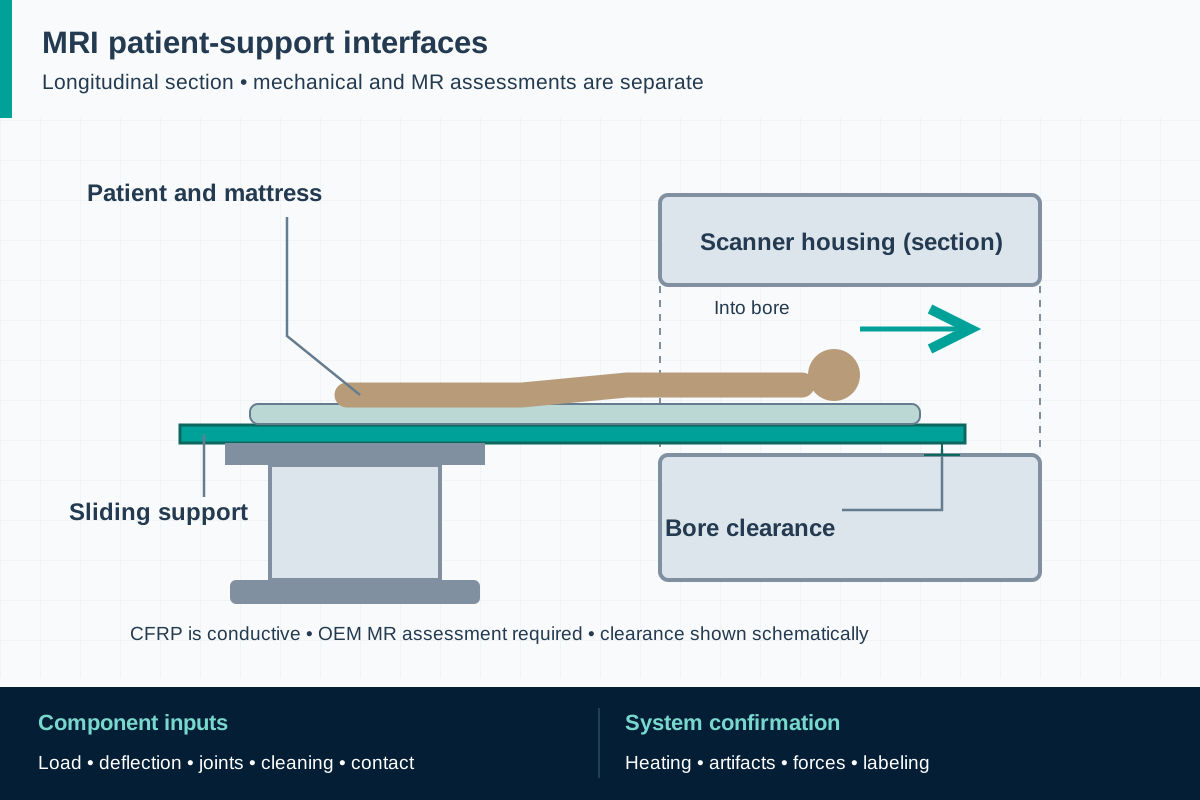
FIG. 1 Composite medical support profile componentsComponent schematic · not to scale
Longitudinal section: the patient and sliding top travel along the open bore. The clearance bracket marks the gap below the support; conductive CFRP and connected hardware require OEM MR assessment.

Component scope and material selection

F1 can assess a component qualification inquiry for straight composite profiles and potential drawing-based processing. This is not a catalog of cleared medical devices or MR-labeled replacement parts, and no F1 medical certification is asserted.

Select the complete material system

Candidate GFRP or CFRP thermoset construction with a controlled composition and finish, subject to the OEM's mechanical, biological and MR evaluation where applicable.

Components to specify

  • Straight longitudinal support blanks
  • Custom equipment rail sections
  • Cut and proposed machined profiles to a controlled OEM drawing

Start with the device manufacturer's drawing

Identify the component's intended use, delivery state and relation to the finished device. A profile blank, a covered internal rail and an independently usable accessory have different specification and regulatory questions. Include the responsible OEM, market and qualification contact in the inquiry so that material development does not proceed against an undefined medical requirement.

Provide the functional envelope, support locations, loads, assembly interfaces and datum scheme. State which surfaces need machining and which must remain protected. If a design requires a curved shell, local thickening or complex inserts, review the manufacturing route instead of assuming that continuous pultrusion produces the entire finished shape.

ReferencesRöchling Industrial: life-sciences applicationsRöchling Industrial: pultrusion process and material selectionRegulation (EU) 2017/745: Medical Device Regulation

Freeze composition beyond the fiber name

Record the proposed reinforcement, resin, additives, color, surface layer and any adhesive, insert or coating within the supplier's delivery scope. Agree on what composition information the OEM requires for assessment and how it will be provided. A trade description such as carbon composite or medical plastic is not a complete material specification.

Select glass or carbon construction against the defined requirements and evaluation plan. Electrical conductivity and magnetic behavior matter when the component is associated with an MR environment; stiffness and section size alone cannot settle the choice. An accepted formulation should have a defined change-notification process, including apparently minor pigment or surface-finish substitutions.

ReferencesRöchling Industrial: pultrusion process and material selectionElectrical properties of carbon-fiber support systemsFDA MR guidance: full October 2023 document

Define edges, surfaces and assembly interfaces

Specify edge condition, allowed surface defects, finish, cleanliness and protection during delivery. Machined holes and ends can expose fibers or create liquid-retaining details, so evaluate the actual finished condition. Coordinate covers, inserts and load-spreading faces with the device designer and identify surfaces requiring inspection before assembly.

List the expected cleaning exposures and whether the delivered component can contact a patient directly or indirectly. The OEM determines the biological evaluation and any material or process evidence needed. Neither a generic resin datasheet nor a favorable result on an unfinished sample covers every finished surface, adhesive or cleaning history.

ReferencesISO 10993-1:2025: biological-safety evaluationISO 14971:2019: medical-device risk management

Agree on the component verification package

Before making prototypes, identify the required dimensional, mechanical and environmental evidence and the configuration represented by each test. For an integrated scanner-table part, use the OEM's equipment assessment and applicable IEC 60601-2-33 route; FDA's 2023 MR guidance excludes the MR system and associated components. A separate device or accessory requires its own scope review. Material screening does not independently authorize an MR label on the final support.

Trace each accepted result to the drawing, material and manufacturing revision. Separate supplier component release from OEM device release. If the customer needs audited medical supplier controls, specialized metrology or third-party testing, confirm those requirements and responsibilities before treating the inquiry as a production order.

ReferencesFDA: testing and labeling medical devices in the MR environmentIEC 60601-2-33:2022: diagnostic MR equipmentFDA: Quality Management System Regulation

Deliver traceable parts and preserve their configuration

Agree on lot identification, inspection records, packaging, shelf or storage conditions where relevant, and the handling of nonconforming parts. Packaging should protect critical surfaces and keep parts associated with their documentation. Marking must be legible without creating a new structural defect or contaminating a contact or bond surface.

Require an OEM review route for material, process and interface changes after qualification. Define the records needed for later replacement and whether service modifications are allowed. QMSR and EU medical-device obligations depend on the actual supplied item and legal role; component traceability supports the OEM's work but does not substitute for the required device conformity assessment.

ReferencesFDA: Quality Management System RegulationRegulation (EU) 2017/745: Medical Device RegulationISO 14971:2019: medical-device risk management

Standards, regulations and acceptance evidence

The project authority determines the governing jurisdiction and adopted edition. The scope of each reference matters: a material test, an equipment requirement and a workplace rule answer different questions.

Guidance · United States medical-device regulatory submissions and labeling

FDA MR testing and labeling guidance, October 2023

MR safety, compatibility and labeling recommendations for medical devices within its stated scope, including independently supplied devices or accessories where applicable.

Application boundary: Excludes the MR system and associated components. Integrated scanner-table rails follow the OEM's equipment qualification, including the applicable IEC 60601-2-33 route. For those parts this nonbinding guidance provides hazard and terminology background, not a direct component approval or labeling route.

ReferencesFDA: testing and labeling medical devices in the MR environmentFDA MR guidance: full October 2023 documentIEC 60601-2-33:2022: diagnostic MR equipment

Standard · Items entering MR environments where this marking practice is adopted

ASTM F2503-26

MR marking terminology, icons and information accompanying medical devices and other items.

Application boundary: The active publisher edition is 2026; regulatory recognition may identify a different edition. Mandatory provisions do not cover image artifacts. The label must follow a supported assessment of the item, not just its resin or reinforcement name.

ReferencesASTM F2503-26: marking items for the MR environment

Standard · Diagnostic MR equipment programs using the IEC route or an applicable national adoption

IEC 60601-2-33:2022

Particular basic-safety and essential-performance requirements for MR diagnostic equipment; the publisher's corrected copy includes interpretation sheets from 2023 and 2025.

Application boundary: Equipment-level scope. A profile certificate cannot establish compliance of the patient-support mechanism, scanner or accessory. Confirm the contractual and regulatory edition with the OEM.

ReferencesIEC 60601-2-33:2022: diagnostic MR equipment

Standard · Medical-device development and biological evaluation where applicable

ISO 14971:2019 and ISO 10993-1:2025

Device risk management and a risk-based evaluation of biological safety, informed by the nature and duration of body contact.

Application boundary: A structural support hidden behind a cover and a directly contacted surface may need different evidence. A resin ingredient report is not a blanket finished-component biological evaluation. Check the regulator's recognized edition as well as the publisher's current edition.

ReferencesISO 14971:2019: medical-device risk managementISO 10993-1:2025: biological-safety evaluation

Regulation · United States; applicable finished-device manufacturers

21 CFR Part 820: QMSR

The QMSR has applied since 2 February 2026 and incorporates ISO 13485:2016, with FDA-specific provisions.

Application boundary: A supplier of an unfinished structural profile is not automatically a finished-device manufacturer. Selling a functioning device or accessory changes the analysis; supplier controls still need agreement with the OEM. No F1 medical QMS certification is asserted here.

ReferencesFDA: Quality Management System Regulation

Regulation · European Union medical devices and accessories within scope

Regulation (EU) 2017/745

The applicable device manufacturer's obligations, risk management, technical documentation and general safety and performance requirements.

Application boundary: An OEM drawing or intended hospital use alone does not establish a component's regulatory classification or CE status. Determine the actual supplied item and intended purpose using the current consolidated regulation; a material supplier's report does not replace device conformity assessment.

ReferencesRegulation (EU) 2017/745: Medical Device Regulation

Public source scopes were reviewed on . These references do not establish certification of an F1 product. Agree the offered grade, finished component, test specimen and acceptance criteria in the project specification.

Composite medical support profile components: practical questions

Can these profiles be ordered as MR Safe stock material?

No such stock designation is claimed. Any MR-related designation must follow a supported assessment of the actual item and its configuration by the responsible device team.

Is ISO 13485 certification claimed for F1 on this page?

No. Any required medical quality-system qualification and supplier audit must be reviewed as part of the inquiry before a supply commitment.

Can a profile of the same size replace an existing scanner-table rail?

Dimensional similarity is insufficient. Replacement requires the OEM's approval of material, mechanical behavior, interfaces, cleaning compatibility and relevant MR or device evidence.

Technical sources and further reading

Industry sources establish where the application is used. Authority sources define the published scope of standards and regulations. The design and procurement checklists above are an original engineering synthesis for project review.
  1. Industry application evidence

    Röchling Industrial: life-sciences applications

    The patient-stretcher example explicitly identifies a pultruded composite profile used in an MRI scanning system. It is application evidence, not F1 medical experience or evidence that all listed material combinations have identical MR behavior.

  2. Industry application evidence

    Röchling Industrial: diagnostic-equipment components

    Identifies patient beds, coil supports and machined components. The page includes multiple materials and manufacturing processes; not every illustrated part is pultruded.

  3. Industry application evidence

    Röchling Industrial: pultrusion process and material selection

    Explains how fiber architecture, resin and section affect profile properties. It does not establish clinical, MR or biological suitability.

  4. Standard or public authority

    FDA: testing and labeling medical devices in the MR environment

    October 2023 nonbinding guidance for in-scope devices entering an MR environment. The full document excludes the MR system and its associated components; it is terminology and hazard background for integrated scanner-table parts, not their direct qualification route.

  5. Standard or public authority

    FDA MR guidance: full October 2023 document

    Page 2 excludes MR systems and associated components, giving spacing pads and coils as examples. Direct application is limited to devices within scope. Integrated scanner supports follow the OEM's equipment assessment; the document provides relevant MR terminology and hazard background.

  6. Standard or public authority

    ASTM F2503-26: marking items for the MR environment

    Active 2026 edition for MR marking and associated information. Image artifacts are outside its mandatory scope; a marking practice does not itself qualify a supplied material.

  7. Standard or public authority

    FDA: MRI safety information for professionals

    Describes RF-related burns as well as patient-table pinch, crush and fall hazards. Useful context for defining the complete support assembly's risks.

  8. Standard or public authority

    IEC 60601-2-33:2022: diagnostic MR equipment

    Edition 4; publisher's corrected November 2025 copy incorporates the 2023 and 2025 interpretation sheets. Equipment basic safety and essential performance, not certification of an isolated profile.

  9. Standard or public authority

    ISO 14971:2019: medical-device risk management

    Official scope for medical-device risk management through the life cycle. The device manufacturer determines risks and acceptance; this page makes no F1 conformity claim.

  10. Standard or public authority

    ISO 10993-1:2025: biological-safety evaluation

    Current sixth edition replaces 2018. Contact nature and exposure inform a risk-based biological evaluation. Publication and regulatory recognition are separate questions.

  11. Standard or public authority

    FDA: Quality Management System Regulation

    Effective 2 February 2026; 21 CFR Part 820 incorporates ISO 13485:2016. FDA explains applicability to finished-device manufacturers and the definition including accessories.

  12. Standard or public authority

    Regulation (EU) 2017/745: Medical Device Regulation

    Official legal entry, including Article 10 and Annex I. Follow the current consolidated text for the project; the manufacturer's obligations depend on the actual device or accessory and intended purpose.

  13. Standard or public authority

    Electrical properties of carbon-fiber support systems

    Primary experimental research showing that carbon-fiber structural systems have important electrical behavior at high frequency. Not an MR safety test or medical-device approval.

  14. Standard or public authority

    AAPM Task Group 334: accessories in the MR environment

    Professional task-group guidance for radiotherapy immobilization devices and accessories in MR environments. Used for configuration and compatibility considerations, not as a universal patient-table specification.

Prepare a component qualification brief

Provide the application and acceptance requirements so we can assess material, geometry, sampling and the proposed supply scope. We reply within one business day; a formal quotation follows once the specification and delivery are reviewed.

01 · OEM drawing, revision, intended use, market and delivery boundary.
02 · Profile geometry, datums, tolerances, machined features and critical surfaces.
03 · Load cases, interface loads, life and permissible deformation.
04 · Proposed composition, finish and excluded or controlled additives and hardware.
05 · Contact scenario, cleaning exposures and relevant assessment responsibilities.
06 · MR-related location, connected configuration and required device-team evaluation.
07 · Prototype and production quantities, documentation, supplier controls and acceptance authority.
Get a quoteWhatsApp